GTO suite
57 opssequence · format · genomic · amino acid · info · text
Workflows
Drag tools into a workflow and test it in your browser on small data. Export it as one file, ready to run where your data lives.
How it works
Drag tools onto the canvas and connect them. Each connection shows the file type it carries.
Run it in your browser on a small sample. Results open next to the graph, and nothing is uploaded.
Export the workflow as one JSON file, then share it or run it where the data is.
Where it can run ↓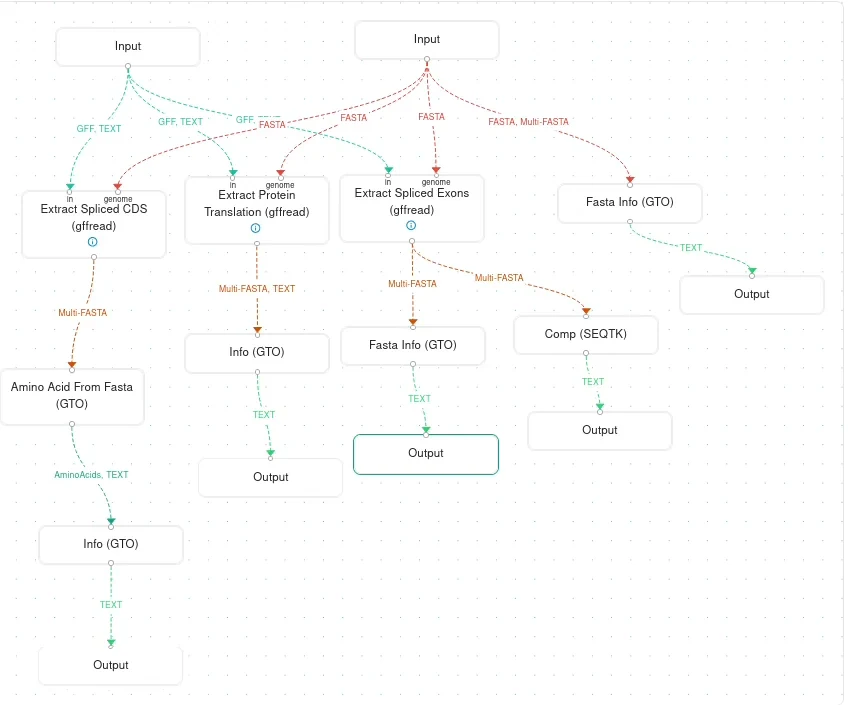
See it in action
Tutorials
14 hands-on tutorials with small example data and the result you should get. Each one runs in your browser and ends with an exercise. New here? Start with “Map reads to a reference”.
One input, one tool: the simplest workflow.
Each tool feeds the next.
Align reads with minimap2, sort them with samtools and count how many mapped. Every move in the editor is shown.
minimap2 · samtools
Call a sample’s consensus with ViralConsensus and check that it finds the mutations you know are there.
minimap2 · samtools · ViralConsensus
Trim primers, call variants and build a consensus with iVar, as in SARS-CoV-2 surveillance.
minimap2 · samtools · iVar
Put a phage genome back together without a reference, then check it against the known genome.
fermi-lite · minimap2
Align eight sequences with muscle, infer their tree with FastTree and compare them with TN93.
muscle · FastTree · TN93
One input feeds several tools, so you compare results side by side.
Count bases, cut a gene out of a genome and read it from the other strand.
seqtk
Filter low-quality reads with fastp and count what is left, before and after.
fastp · seqtk
Measure how much of a genome is covered and list the regions where an amplicon failed.
minimap2 · samtools · gawk
Summarise a VCF, keep the variants with enough quality and depth, and turn them into a table.
bcftools
Two or more files meet in one tool.
Filter read pairs and merge the ones that overlap into single longer reads.
fastp · seqtk
Overlap variants with gene coordinates and see why one variant can land in two genes.
bcftools · bedtools
Align two phage genomes, measure how similar they are and see why a circle aligns in two pieces.
minimap2
No prior knowledge needed; good for a class.
Checking and preparing sequencing data.
Filter low-quality reads with fastp and count what is left, before and after.
fastp · seqtk
Filter read pairs and merge the ones that overlap into single longer reads.
fastp · seqtk
Measure how much of a genome is covered and list the regions where an amplicon failed.
minimap2 · samtools · gawk
Viral genomes and genomic surveillance.
Call a sample’s consensus with ViralConsensus and check that it finds the mutations you know are there.
minimap2 · samtools · ViralConsensus
Trim primers, call variants and build a consensus with iVar, as in SARS-CoV-2 surveillance.
minimap2 · samtools · iVar
Working with variants.
Genes, assemblies and genome comparison.
Predict a phage’s genes with Prodigal, act on its error message and compare with the annotation.
Prodigal
Put a phage genome back together without a reference, then check it against the known genome.
fermi-lite · minimap2
Align two phage genomes, measure how similar they are and see why a circle aligns in two pieces.
minimap2
How sequences and species are related.
Raw sequencing reads.
Reads placed on a reference.
Align reads with minimap2, sort them with samtools and count how many mapped. Every move in the editor is shown.
minimap2 · samtools
Measure how much of a genome is covered and list the regions where an amplicon failed.
minimap2 · samtools · gawk
Where a sample differs from the reference.
Call a sample’s consensus with ViralConsensus and check that it finds the mutations you know are there.
minimap2 · samtools · ViralConsensus
Trim primers, call variants and build a consensus with iVar, as in SARS-CoV-2 surveillance.
minimap2 · samtools · iVar
Summarise a VCF, keep the variants with enough quality and depth, and turn them into a table.
bcftools
Overlap variants with gene coordinates and see why one variant can land in two genes.
bcftools · bedtools
Whole sequences, their genes and assemblies.
Count bases, cut a gene out of a genome and read it from the other strand.
seqtk
Predict a phage’s genes with Prodigal, act on its error message and compare with the annotation.
Prodigal
Put a phage genome back together without a reference, then check it against the known genome.
fermi-lite · minimap2
Align two phage genomes, measure how similar they are and see why a circle aligns in two pieces.
minimap2
Alignments and trees.
Teaching a class? Each tutorial includes its data and a finished workflow file, so students can import it and catch up at any step.
Catalogue
The 34 signed recipes in the public BioChef catalogue, grouped by what they do. Every tool runs as WebAssembly in the browser; 15 also have a native build for the agent.
sequence · format · genomic · amino acid · info · text
minimap2 · lastz · mummer4/nucmer · ssw · ksw2 · edlib
bcftools · ivar · viralconsensus
samtools view/sort/stats · fastp · samblaster
muscle · kalign · abPOA · fasttree · ASTER · tn93
bedtools intersect · cgranges coverage
fermi-lite assemble · prodigal · modbam2bed
seqtk · gffread — subseq · convert · filter
gawk · grep · sed · jq · coreutils
trimal · wgsim · yak
Badges count recipes per family (GTO counts operations); 188 operations in all. New recipes are added by the community. Workflows can pipe FASTA, FASTQ, BAM, SAM, CRAM, VCF, BCF, BED, GFF, JSON, LIST and more between tools.See the full catalogue in the app →
Runtime
Live
For building and testing. Tools run client-side as WebAssembly on small data, with nothing to install and nothing uploaded.
Emscripten · biowasm · signed digest from the registry
In development
For full runs. The BioChef agent runs the same signed tools on your workstation, a server or a secure environment, so large or sensitive data never has to move.